


Welcome to the website of Anh Khoa Augustin Lu !
I am currently a senior researcher at the National Institute for Materials Science.
You can find information about my research activities on this website.
| Name | Anh Khoa Augustin Lu |
| Country | Belgium  |
| Position | Senior researcher |
| Research group | Research Center for Materials Nanoarchitectonics (WPI-MANA), Quantum Materials Simulation Group |
| Hobbies | Piano, beach tennis, football |
| Favourite dishes | Phở bò, tonkatsu |

My research focuses on the relation between the atomic structure and the properties of materials. It combines first-principles calculations, molecular dynamics with machine-learning interatomic potentials, and structural analysis using Voronoi tessellation, the p3 code, unsupervised learning (TS-LPP), and topological data analysis (differential persistence diagrams), with the aim of understanding these materials and designing new ones. The materials studied are semiconductors and oxides, metallic glasses, and two-dimensional materials.
Oxides and semiconductors are central to electronic and energy devices, where defects, interfaces and external fields control their behavior. Topics include ion transport in ZrO2 under an electric field, the structure of silica under pressure, and semiconductor/oxide interfaces such as Si/SiO2.

Metallic glasses can display outstanding mechanical properties, yet their atomic structure remains poorly understood. Unlike crystals, they lack long-range order, so the link between their atomic configurations and their macroscopic properties is still an open question. Atomistic simulations are used to study their short- and medium-range order and its relation to crystallization, shear-induced segregation, and strength.

Since graphene, two-dimensional materials have grown into a large family, from transition metal dichalcogenides to Xenes. First-principles calculations are used to study doping and defects in transition metal dichalcogenides such as Nb-doped WSe2, new bilayer GaN structures, and magnetism in Cr2Ge2Te6, with applications in nanoelectronics and spintronics in view.

To describe structure beyond the first-neighbor shell, locally averaged atomic fingerprints are combined with dimensionality reduction such as two-step locality preserving projections (TS-LPP). This reveals subtle structural differences and the medium-range order of disordered materials such as silica.
To quantify subtle structural changes in disordered systems across multiple length scales, persistent homology is extended with differential persistence diagrams (DPDs). By directly contrasting topological features—such as rings and cavities—between structures prepared under different conditions, DPDs isolate preparation-dependent short- and medium-range order in amorphous materials such as vapor-deposited Cu-Zr metallic glasses.
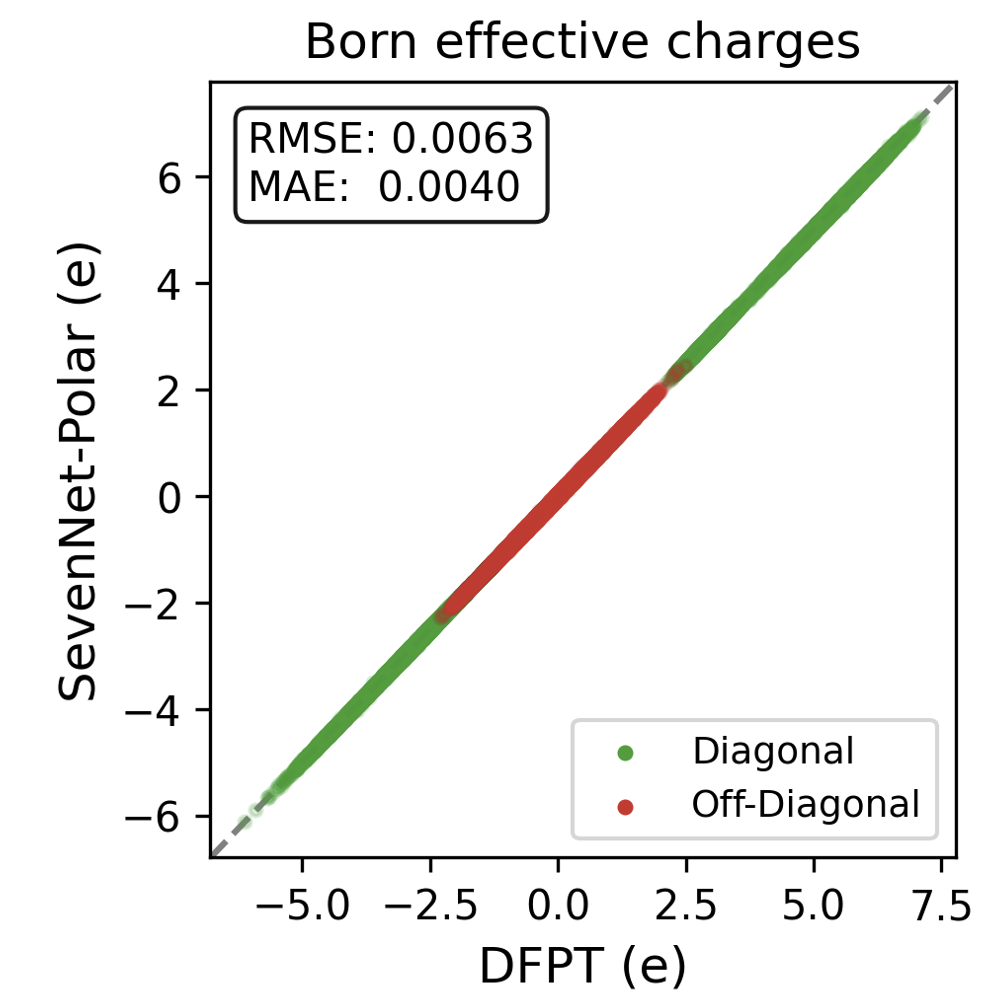
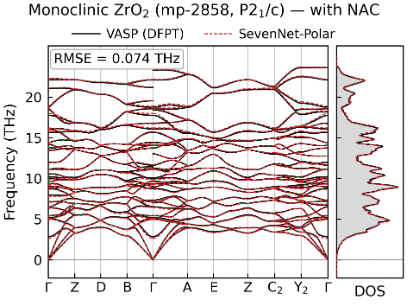
Simulating materials under an electric field requires an accurate description of the forces induced by that field, captured here through Born effective charge tensors. SevenNet-Polar, an equivariant graph neural network built on SevenNet, predicts Born effective charges together with energies, forces and stress, in a single multitask model. Its predictions closely match first-principles calculations for all four quantities, which makes large-scale molecular dynamics under an applied electric field possible. The same model also gives phonon band structures with non-analytical corrections.
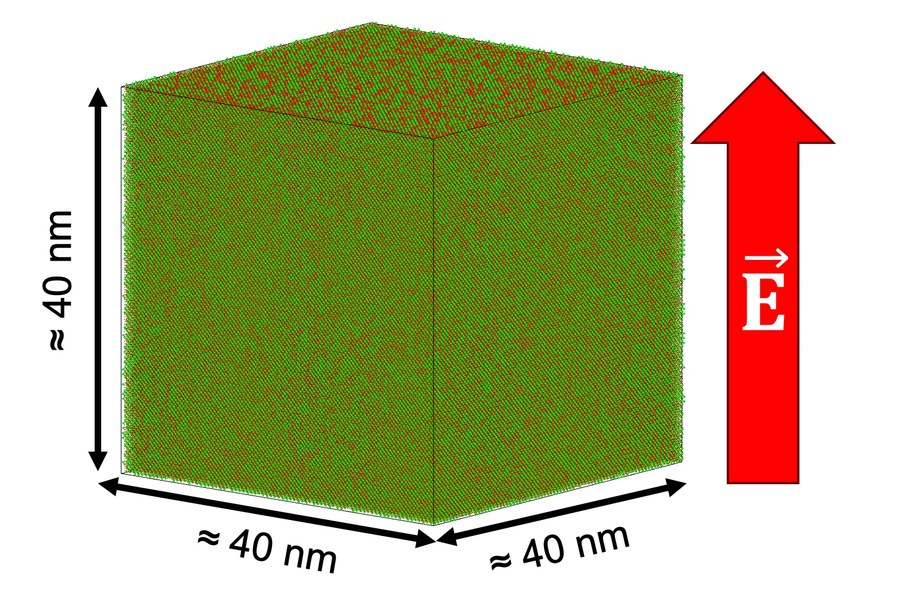
Contributions to the development of CONQUEST, a linear-scaling DFT code for first-principles calculations on very large systems. With SevenNet-Polar, molecular dynamics of more than five million atoms under an electric field were run on the full GPU partition of the Wisteria/BDEC-01 supercomputer, as part of the University of Tokyo Large-Scale HPC Challenge 2026.
Research Center for Materials Nanoarchitectonics (WPI-MANA)
Quantum Materials Simulation Group
Interested in a collaboration? Feel free to contact me:
Anh Khoa Augustin Lu
NIMS
Namiki 1-1, Tsukuba, Ibaraki 305-0044
Japan
e-mail: lu[dot]augustin[@t]nims.go.jp


